Key takeaways
Refunding a customer does not close the safety assessment; assign someone to handle it.
EU serious undesirable effects and US serious adverse events have different definitions, duty holders and reporting clocks.
Assess serious-event reporting and the need for recall separately, without making either wait for the other.
A recall procedure needs distribution records, recipient follow-up, quantity reconciliation and corrective actions.
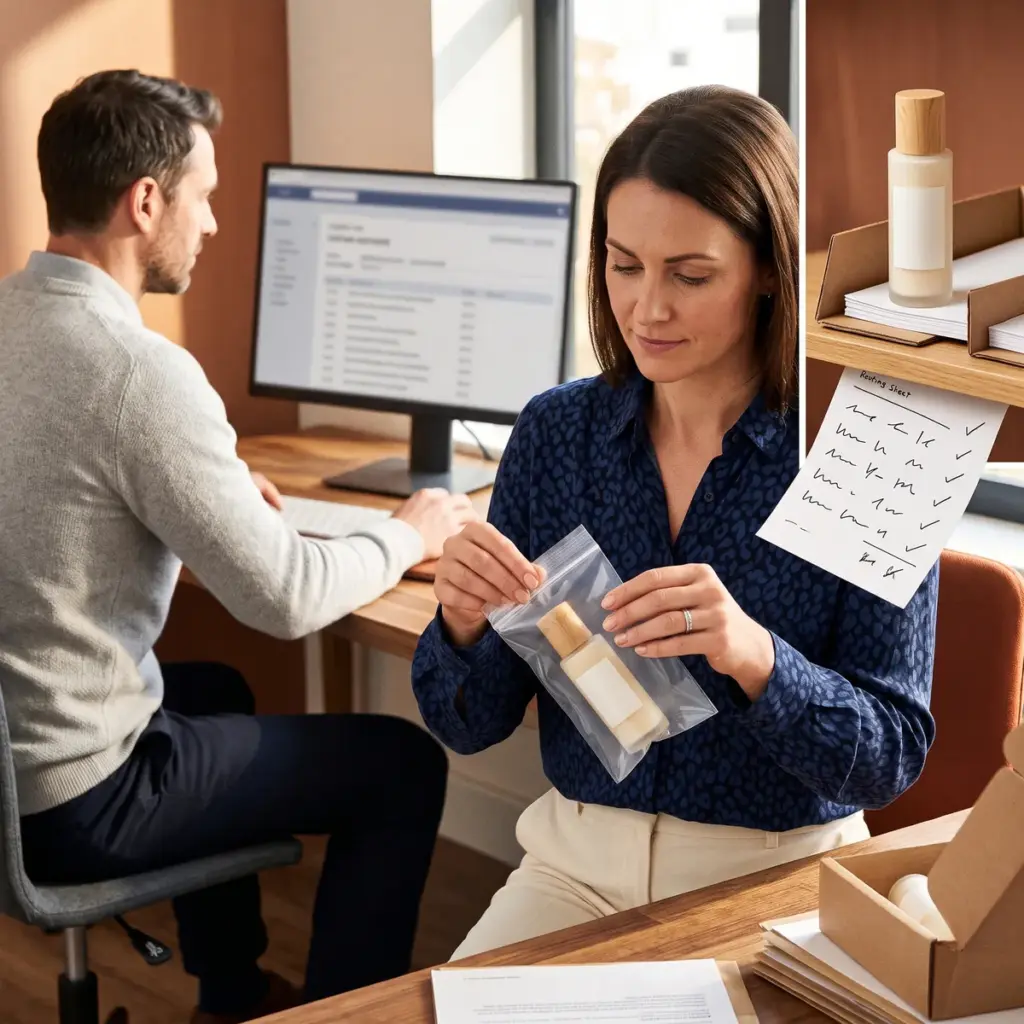
A refunded complaint may still need a safety investigation. Customer care handles the purchase problem; quality identifies the product and investigates, while regulatory staff assess reporting duties. Assign owners for both the refund and the safety assessment.
Brands selling in the EU and US can use one complaint intake system, provided staff assess each market’s reporting duties separately. The definitions and reporting deadlines differ. The procedure also needs batch records that let quality trace the affected stock. This guide distinguishes recommended company procedures from the legal duties cited below.
Give every complaint a traceable record
Assign a case number and preserve the customer’s account without turning it into a diagnosis. Record receipt date and time, country of use, product name and size, batch code if available, purchase channel, use dates, reported problem and a safe way to follow up. Save the relevant label version and photographs where available and appropriate.
For a reported health effect, seek information on onset, duration, outcome and medical attention, with suitable privacy controls. Record missing information as missing. Staff should not dismiss a case because a customer cannot find the batch code, and should never ask the customer to use the product again to reproduce a reaction. For a reaction, FDA advises stopping use and contacting a healthcare provider.
Train retail, social, sales and customer-care teams to forward health-related reports promptly. Define internal escalation times and deputies; these are company controls, not substitute legal deadlines. Keep medical information accessible only to authorised staff, and separate regulatory follow-up from refund approval.
Assess EU and US reportability separately
Staff need to recognise potential serious events and refer them promptly for specialist assessment. Reportability depends on the defined outcomes, rather than how strongly a customer describes a reaction.
| Market | Duty holder and trigger | Reporting requirement |
|---|---|---|
| EU | Responsible person and distributors: serious undesirable effects known to them or reasonably expected to be known | Notify the competent authority where the effect occurred without delay under Article 23; include product identification and corrective measures, if any |
| US | Responsible person: receipt of a report of a serious adverse event associated with cosmetic use in the US | Submit to FDA no later than 15 business days after receipt, with the label on or within the retail packaging |
| US follow-up | Responsible person: new and material medical information received within one year of the initial FDA report | Submit no later than 15 business days after receipt of that information |
Under EU Regulation 1223/2009, Article 2, a serious undesirable effect is an undesirable effect resulting in temporary or permanent functional incapacity, disability, hospitalisation, congenital anomalies, an immediate vital risk or death. Article 23 sets the duty as “without delay”. The Commission’s SUE reporting guidelines, section 2.4.3, interpret the timeframe as within 20 calendar days from awareness by any employee, whatever their role. Do not treat that guidance period as permission to postpone notification.
The EU guidance gives minimum information for an initial notification: an identifiable reporter, the nature and onset date of the alleged SUE, and a specifically identifiable product. It allows follow-up reports and says all SUE cases should be notified except those whose causality is assessed as excluded within the timeframe. Route a potential case to qualified staff promptly rather than waiting for a completed laboratory investigation.
The US definition published by FDA differs. It includes death, a life-threatening experience, inpatient hospitalisation, persistent or significant disability or incapacity, congenital anomaly or birth defect, infection, and specified significant disfigurement, as well as medical or surgical intervention needed, on reasonable medical judgment, to prevent those outcomes. The responsible person is the manufacturer, packer or distributor whose name appears on the label, not automatically the contract factory or EU responsible person.
The US reporting deadline runs from the responsible person’s receipt of the report, without waiting for causality testing. 21 USC 364a establishes the deadlines in the table. FDA’s current instructions also call for medical or other follow-up information received within a year to be submitted within 15 business days. Confirm the current submission route on FDA’s complaint-reporting page; consumer voluntary reporting does not replace the responsible person’s mandatory report. Use the MoCRA requirements guide for the wider US workflow.
Investigate without losing the original evidence
Quality should check the batch record, release results, deviations, ingredient lots, packaging version and distribution history. Look for complaints involving the same batch or failure mode, checking for duplicate reports of one event. Compare the evidence from opened returns and unopened retained samples without assuming either represents every unit.
Agree sample handling before arranging a return. Photograph the condition, record who received it and when, protect any seals, and document storage and transfers. Have a competent laboratory define appropriate sampling and methods. Contamination in an opened return does not by itself establish contamination across the batch. Interpret the result alongside manufacturing records and the other evidence.
Use the batch-release and certificate-of-analysis guide to identify the release evidence worth retrieving. If the investigation points to preservation, distinguish product microbial quality from preservative efficacy using the challenge-test guide.
Decide whether to hold stock, withdraw or recall
An internal stock hold prevents further shipment while the team assesses a concern. In EU terminology, withdrawal prevents supply-chain availability, while recall seeks return of product already supplied to end users. Define those actions in the procedure so a warehouse stop is not mistaken for a consumer recall.
EU Articles 5 and 6 impose corrective-action duties on responsible persons and distributors when they consider or have reason to believe a product is non-compliant, with withdrawal or recall as appropriate. Where the product presents a human-health risk, the responsible person must immediately inform authorities in the Member States where it made the product available and where the PIF is accessible. Distributors must immediately inform the responsible person and authorities where they made it available. Authorities can require measures under Articles 25 and 26; Article 27 addresses provisional measures for serious risk even where listed requirements are met. These duties can require a company to act before an authority orders it to do so.
In the US, FDA says most cosmetics recalls are voluntary, whether initiated by the firm or requested by FDA. MoCRA also gives FDA mandatory recall authority where there is a reasonable probability that a cosmetic is adulterated or misbranded and exposure will cause serious adverse health consequences or death, if the responsible person fails to act voluntarily when given the opportunity. FDA announced final guidance on this authority on 22 October 2026.
Assess the reporting duty separately from the need for a recall. A report does not itself establish product-wide risk, and a quality defect may require action before any injury is reported.
Prepare the recall file before contacting customers
Give a named incident lead authority to coordinate quality, regulatory, operations and customer communications. Log which products and batches are affected, the reasons for that scope and any uncertainty. Poor batch traceability may require a wider scope; it does not establish that unidentified units are unaffected.
The recall checklist should cover:
- Product names, pack images, batch identifiers, markets and reasons for action.
- Stock and shipment blocks covering warehouses, distributors and online fulfilment.
- A distribution list and approved retailer and consumer instructions, including returns or disposal.
- Required authority contacts, report owners and submission records.
- Checks that recipients received and acted on the notice, with follow-up for nonresponders.
- Reconciliation of affected quantities and documented disposition, without double-counting returned stock.
FDA’s voluntary-recall policy recommends notifying the relevant FDA office, status reports and effectiveness checks; coordinate the strategy and termination with FDA as applicable. EU contacts and measures must follow the relevant authority’s instructions. Do not declare completion solely because the brand has issued refunds.
Hypothetical example: complaints about an unusual odour
Suppose customer care receives odour complaints about one lotion batch, with no health effect initially reported. Quality blocks remaining stock and arranges controlled examination of retained and returned units while regulatory staff assess the facts. The team checks adjacent batches and shared raw-material lots rather than assuming the printed batch code defines the entire problem.
If a later report describes an infection associated with use in the US, the responsible person promptly assesses reportability and tracks the deadline from receipt of that report, independently of the recall decision. Evidence of a wider safety problem would also inform which products to include in the action and what to communicate to authorities.
Complete corrective actions and test the procedure
Address the confirmed cause through corrective actions. If the cause remains uncertain, record that uncertainty and the safeguards the evidence supports. Update the relevant safety records; EU undesirable-effect information can affect the PIF and safety report. Keep US adverse-event records for the statutory period: generally six years, or three for qualifying small-business responsible persons under the conditions in section 364a(e).
Run a mock recall using a real batch and distribution records, without sending customer notices. Check whether staff can locate stock, reach decision-makers and reconcile quantities. Revise any steps that failed during the exercise and repeat the affected checks.
Sources and references


Frequently asked questions
Can a brand wait for laboratory results before submitting a US serious adverse event report?
Can a brand wait for laboratory results before submitting a US serious adverse event report?
No. The statutory deadline is no later than 15 business days after the responsible person receives the serious adverse event report, regardless of whether the laboratory investigation is complete. Submit as required and send subsequent information through the follow-up process.
Does an EU distributor have its own SUE reporting duty?
Does an EU distributor have its own SUE reporting duty?
Yes. Article 23 places the notification duty on responsible persons and distributors. The distributor can collaborate with the responsible person, but that arrangement does not remove the distributor’s legal duty or justify delay.
Does one serious report automatically require a recall?
Does one serious report automatically require a recall?
No. Assess the event-reporting duty and the product risk separately. A report alone does not establish the need for recall, while a serious product defect can require corrective action even without a reported injury.